
FDA更新510(k)豁免指南,豁免的低风险医疗器械能直接进入美国市场吗?

FDA更新510(k)豁免指南,豁免的低风险医疗器械能直接进入美国市场吗?
2026年6月5日,美国食品药品监督管理局(FDA)更新了《Intent to Exempt Certain Unclassified Medical Devices from Premarket Notification Requirements》指南,涉及部分未分类医疗器械的510(k)上市前通知豁免事项。FDA表示,对于指南中列出的相关产品,在正式豁免生效前,现阶段不打算强制执行其510(k)要求。
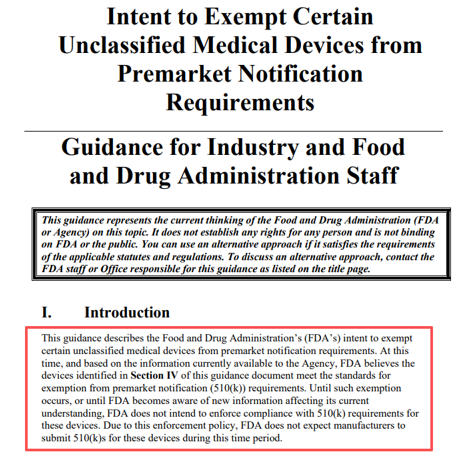
Intent to Exempt Certain Unclassified Medical Devices from Premarket Notification Requirements
Guidance for Industry and Food and Drug Administration Staff
This guidance represents the current thinking of the Food and Drug Administration (FDA or Agency) on this topic. It does not establish any rights for any person and is not binding on FDA or the public. You can use an alternative approach if it satisfies the requirements of the applicable statutes and regulations. To discuss an alternative approach, contact the FDA staff or Office responsible for this guidance as listed on the title page.
I. Introduction
This guidance describes the Food and Drug Administration's (FDA's) intent to exempt certain unclassified medical devices from premarket notification requirements. At this time, and based on the information currently available to the Agency, FDA believes the devices identified in Section IV of this guidance document meet the standards for exemption from premarket notification (510(k)) requirements. Until such exemption occurs, or until FDA becomes aware of new information affecting its current understanding, FDA does not intend to enforce compliance with 510(k) requirements for these devices. Due to this enforcement policy, FDA does not expect manufacturers to submit 510(k)s for these devices during this time period.
这项更新再次引起了企业对医疗器械上市前监管要求的关注。对于计划进入美国市场的企业来说,产品风险等级较低,是否就意味着可以免于510(k)?即使符合豁免条件,还需要办理哪些手续?
低风险医疗器械,都可以免于510(k)吗?
510(k)是医疗器械上市前通知程序。对于需要提交510(k)的产品,企业需要向FDA提供相关资料,证明产品与合法上市的对比器械具有实质等同性。
FDA将医疗器械分为Class I、Class II和Class III三个风险等级。多数Class I产品可以免于510(k),但并非全部如此。企业仍需根据产品分类、对应的法规条款及豁免限制,确认是否需要提交510(k)。
免于510(k)的低风险医疗器械,还需要办理哪些手续?
免于510(k)不等于免除所有FDA监管要求。对于符合豁免条件的产品,企业仍需根据自身业务活动及产品适用的法规,确认以下事项:
企业登记(Establishment Registration)
从事医疗器械生产、进口或其他受监管活动的相关企业,需根据FDA规定确认是否需要进行设施登记,并按要求缴纳年度登记费用。
产品列名(Device Listing)
符合相关要求的企业还需在FDA系统中列名其医疗器械,并提供相应的产品分类代码及活动信息,取得FDA注册编号。
境外企业指定美国代理人(U.S. Agent)
对于需要向FDA登记的境外医疗器械企业,通常还需指定一名美国代理人,协助FDA与境外企业进行沟通。美国代理人并不因此承担制造商的全部监管责任。
赛德斯威如何帮助您完成FDA低风险医疗器械注册?
从产品分类确认到注册申请及后续维护,我们可提供全流程技术支持,包括:
●FDA医疗器械分类及适用要求确认
●510(k)豁免条件核对
●FDA企业登记(Establishment Registration)
●医疗器械产品列名(Device Listing)
●美国代理人相关服务
●FDA注册资料准备及申请支持
赛德斯威(SATISFY)可为医疗器械企业提供FDA注册咨询与申请支持,协助企业确认产品适用的监管要求。详情请联系赛德斯威客服。